
Is Obesity Genetic or Behavioral? How Epigenetics Is Rewriting the Story of Body Weight
Quick Answer
Watch Related Video
A pregnant mouse was exposed to bisphenol A (BPA), a chemical used to make polycarbonate plastic. Her descendants became obese, and the effect was transmitted all the way to the sixth generation (Jung et al., Proceedings of the National Academy of Sciences, 2022). None of those later generations were exposed to the chemical themselves. No gene was mutated. What changed was the way their DNA was packaged and read.
If you still believe obesity comes down to laziness or a lack of willpower, that experiment should give you pause. Body weight is shaped not only by what you eat, but by chemical marks layered on top of your genes. Some of those marks were written before you were born. Some may trace back to ancestors you never met. And some, importantly, can still change today.
This is the science of epigenetics, and it is changing how we understand obesity as a chronic disease.
What Is Epigenetics? The Hardware and the Software
Every cell in your body carries the same DNA: the same three billion letters copied into your skin cells, your neurons, and your fat cells. So why does a liver cell behave like a liver cell and a brain cell like a brain cell?
Because DNA is not the whole story.
Think of your DNA as the hardware, the fixed machine you are born with. Epigenetics is the software: the instructions that determine which genes are switched on, which are switched off, and how strongly each one is expressed, without changing a single letter of the underlying code.
Epigenetic changes are stable, sometimes inheritable, changes in gene activity that occur without altering the DNA sequence. They help explain why identical twins, who share the same genome, can grow more different over a lifetime.
The Four Layers of Epigenetic Control
The epigenetic toolkit includes three classic mechanisms (DNA methylation, histone modification, and non-coding RNA) plus a newer fourth layer (Godfrey et al., Lancet Diabetes and Endocrinology, 2017).
- DNA methylation. A tiny chemical tag called a methyl group attaches to a cytosine (C) in the DNA, usually where a C sits next to a G. When these tags cluster around a gene's control switch, they can turn that gene down or off. DNA methylation is chemically stable and can persist for decades, which is why it is the most studied epigenetic mark in obesity research.
- Histone modification. DNA is wound around spool-like proteins called histones. Chemical marks on histone tails control how tightly DNA is wound. Tightly wound DNA is silenced; loosely wound DNA can be read.
- Non-coding RNA. Not all RNA builds proteins. Short RNA molecules can intercept messenger RNA and mark it for destruction, silencing genes before they are ever translated.
- RNA methylation. The same type of methyl tag can be placed on RNA messages themselves, affecting how stable they are and how much protein they produce.
Writers, Readers, and Erasers: Why Diet Reaches Your Genes
These marks are not permanent graffiti. Specialized enzymes write them, erase them, and read them. DNA methyltransferases add methyl tags, while TET enzymes remove them (Lagarde et al., Oncogene, 2024).
Here is the critical link to nutrition. The enzymes that place methyl tags need a methyl donor called S-adenosylmethionine (SAM). SAM is produced through one-carbon metabolism, which depends on folate, vitamin B12, vitamin B6, methionine, choline, and other dietary nutrients (Toraño et al., BioMed Research International, 2016; Donohoe and Bultman, Journal of Cellular Physiology, 2012). Other epigenetic enzymes depend on metabolic fuels such as acetyl-CoA.
In other words, the machinery that controls your genes runs on fuel that comes from your diet and your metabolism. Epigenetics is the physical bridge between your environment and your DNA.
The Agouti Mouse: One Experiment That Changes Everything
Picture two mice. One is round and yellow. The other is lean and brown. They are genetically identical.
The difference comes down to a single switch near a gene called Agouti. When a stretch of DNA upstream of the gene is heavily methylated, the gene stays quiet: the coat is brown and the mouse stays lean. When that stretch loses its methyl tags, the gene is overexpressed. The agouti protein then blocks the melanocortin 4 receptor, a key satiety pathway in the brain, and the mouse develops a yellow coat, obesity, diabetes, and a higher tumor risk (Eichenlaub-Ritter and Pacchierotti, BioMed Research International, 2015; Wolff et al., FASEB Journal, 1998).
What determines where the tags go? The mother's diet during pregnancy.
When pregnant mice were fed extra methyl donors (folic acid, vitamin B12, choline, and betaine), more of their pups were born brown and lean, and the degree of methylation tracked directly with coat color (Wolff et al., FASEB Journal, 1998; Labbé et al., Oncogene, 2015).
Now add BPA. In a landmark study, maternal BPA exposure shifted offspring toward the yellow, obese phenotype by decreasing methylation at the Agouti locus. But when the mother's diet was supplemented with methyl donors such as folic acid, or with the plant compound genistein, the BPA-induced loss of methylation was prevented (Dolinoy et al., Proceedings of the National Academy of Sciences, 2007).
Identical genome. Different maternal environment. Different offspring.
In the interest of scientific honesty, this model has been challenged. A 2021 study found that other variably methylated regions in the mouse genome were largely resistant to maternal BPA and methyl donor exposure, and a later study reported that Agouti coat color was unaffected by maternal BPA (Bertozzi et al., Nature Genetics, 2021; Bourc'his, Nature Genetics, 2021). The agouti mouse remains a powerful teaching model, but it is one model, not the final word.
Four Ways the Environment Writes on Your Genome
The environment does not rewrite your DNA letters. It changes the marks sitting on top of them. Researchers have described several routes.
- Route 1: The supply chain. Methyl tags depend on dietary nutrients that feed SAM production. When methyl donors are scarce, the cell may be unable to maintain methylation, and genes that should be silenced can drift back on (Donohoe and Bultman, Journal of Cellular Physiology, 2012).
- Route 2: Direct interference. Exposures such as air pollution, heavy metals, tobacco smoke, and BPA have been associated with altered DNA methylation patterns.
- Route 3: Hijacking the control panel. Some chemicals, often called obesogens, interact with pathways that regulate fat-cell development, such as PPAR-gamma. Prenatal exposure to low-dose BPA has been linked to faster weight gain, insulin resistance, larger fat cells, and a greater tendency of precursor cells to become fat cells in offspring mice (Zhuang et al., Ecotoxicology and Environmental Safety, 2023). Phthalates such as dibutyl phthalate have shown similar obesogenic effects in animal studies (Li et al., Scientific Reports, 2020).
- Route 4: Chronic stress. Sustained stress signaling through the cortisol system has been associated with epigenetic changes in adipose tissue metabolism (Xiao et al., Nutrition and Metabolism, 2020).
The Showpiece: How a Chemical Can Refold Your Genome
The BPA transgenerational study offers one of the most striking mechanisms in the field. Researchers found that exposing pregnant mice to BPA caused obesity in the grandchildren through increased food intake, and the trait was passed down to the sixth generation. In the sperm of each generation, they found changes in chromatin accessibility at binding sites for a genome-organizing protein called CTCF, located at regulatory elements of the Fto gene, a well-known obesity gene. These elements showed increased contact with two neighboring genes, Irx3 and Irx5, which are involved in developing appetite-controlling neurons (Jung et al., Proceedings of the National Academy of Sciences, 2022).
The decisive experiment: when researchers deleted that single CTCF binding site, the mice ate normally and did not become obese, even when their ancestors had been exposed to BPA. As the authors concluded, epigenetic changes can produce the same phenotypes as genetic variants (Jung et al., Proceedings of the National Academy of Sciences, 2022).
A chemical exposure did not change the DNA sequence. It changed how the genome folded in three dimensions, and that changed appetite across generations.
Gut Bacteria: Another Messenger to Your Genes
When you eat fiber from vegetables, beans, and whole grains, gut bacteria ferment it into short-chain fatty acids. The most potent of these, butyrate, inhibits histone deacetylases, enzymes that tighten DNA packaging. By blocking them, butyrate increases histone acetylation, opens chromatin, and changes gene expression (Mann et al., Nature Reviews Immunology, 2024; Licciardi et al., Antioxidants and Redox Signaling, 2012). Most of this evidence comes from intestinal and immune tissues, so its direct role in human body weight is still being defined (Birigwa et al., Frontiers in Aging, 2026).
The pathway is simple: food feeds bacteria, bacteria make chemical signals, and those signals can influence gene regulation in your cells.
The "Missing Heritability" of Obesity
Obesity clearly runs in families. In a pooled analysis of more than 140,000 twin pairs from 20 countries, the heritability of BMI was about 77% in men and 75% in women in their twenties, declining to about 57% to 59% in older age (Silventoinen et al., American Journal of Clinical Nutrition, 2017).
Yet the genetic variants identified so far, including well-known genes like FTO, explain only a small fraction of that heritability. Scientists call this gap "missing heritability," and no consensus explanation has emerged (Hinney et al., Nature Reviews Endocrinology, 2022). Epigenetics is one of several areas being investigated.
Zooming In: SREBF1, a Fat-Making Switch
SREBF1 is a master regulator of fat production. It drives the conversion of nutrients into fatty acids and triglycerides and helps fat cells develop (Fall et al., Gastroenterology, 2017).
In large human studies, higher methylation at specific sites on SREBF1 is associated with higher triglycerides, lower HDL ("good") cholesterol, central obesity, and type 2 diabetes (Jones et al., Current Atherosclerosis Reports, 2021). Using Mendelian randomization, a genetic technique that helps test cause and effect, researchers linked methylation at one SREBF1 site to BMI, adiposity-related traits, and coronary artery disease (Mendelson et al., PLoS Medicine, 2017). A follow-up analysis estimated that SREBF1 gene expression accounts for roughly 16% to 36% of the relationship between that methylation mark and triglycerides, fasting glucose, and BMI (Keshawarz et al., Scientific Reports, 2023).
But here is the twist: the arrow points both ways. Mendelian randomization also shows that high triglyceride levels themselves drive methylation at this gene (Jones et al., Current Atherosclerosis Reports, 2021). Diet and body fat may help lay down the tag, and once it is there, the tag may feed back into the fat-making program. Cause and consequence, chasing each other in a circle.
The Womb: The Most Important Environment You Can't Remember
The developmental origins of health and disease (DOHaD) framework holds that conditions early in life, especially around conception and during pregnancy, shape lifelong disease risk (Fleming et al., Lancet, 2018).
The Dutch Hunger Winter
In the winter of 1944 to 1945, a German blockade cut off food to the western Netherlands. Daily intake fell to as low as 500 calories (Gomez-Verjan et al., Clinical Genetics, 2020). Decades later, researchers compared people exposed to the famine in the womb with their unexposed same-sex siblings.
Six decades after the famine, people exposed around the time of conception had less DNA methylation at IGF2, an imprinted gene that regulates growth, than their own siblings. The effect was specific to periconceptional exposure, pointing to very early development as a critical window for setting epigenetic marks (Heijmans et al., Proceedings of the National Academy of Sciences, 2008). People prenatally exposed to the famine also had higher rates of obesity, an atherogenic lipid profile, and coronary heart disease (Gomez-Verjan et al., Clinical Genetics, 2020). The absolute methylation differences were small, under 3%, but meaningful relative to normal population variation (Tobi et al., PLoS One, 2012).
Siblings Born Before and After Bariatric Surgery
If scarcity in the womb leaves a mark, what about excess? Bariatric surgery created a remarkable natural experiment.
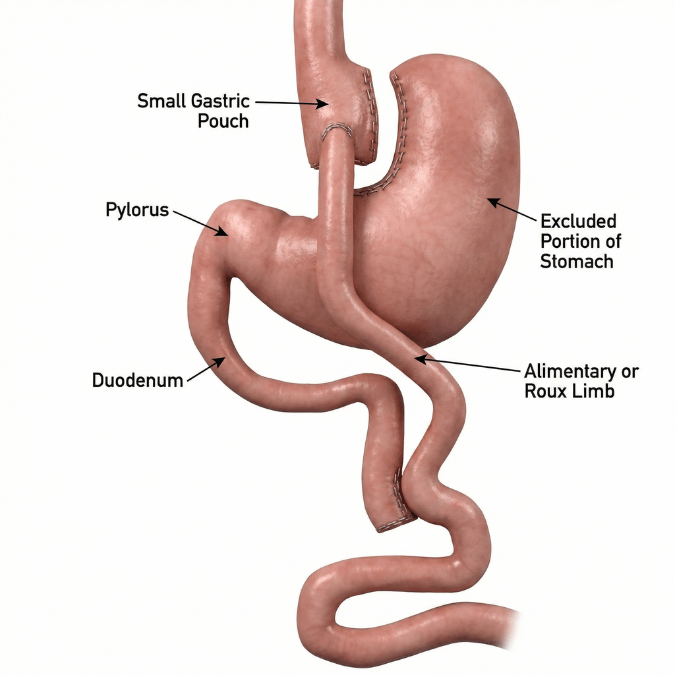
Researchers compared siblings born to the same mother before and after her weight loss surgery. Children born after maternal surgery were less obese and had better cardiometabolic profiles. Across 5,698 genes, methylation differed between the siblings, with a preponderance of genes involved in glucose regulation, inflammation, and vascular disease (Guénard et al., Proceedings of the National Academy of Sciences, 2013). A Swedish study found similar differences in genes involved in insulin signaling, type 2 diabetes, and leptin signaling (Berglind et al., Obesity, 2016).
As the investigators put it, effective treatment of the mother's condition was durably detectable in the methylome of her later children (Guénard et al., Proceedings of the National Academy of Sciences, 2013). A 2023 scoping review cautions that the data remain limited and the effects on offspring weight are inconsistent (Yu et al., Pediatric Research, 2023).
The takeaway is sobering: some of the marks on your genes were shaped by circumstances you never chose. That starting configuration is not your fault.
Beyond the Mother: Grandparents and Fathers
Överkalix, Sweden. Historical harvest records from this remote parish suggested that a boy's food supply just before puberty predicted the mortality of his grandsons, with sex-specific effects along the male line (Alipoor et al., Journal of Internal Medicine, 2025). A replication study nearly 40 times larger, in the Uppsala Multigeneration Study, confirmed that a paternal grandfather's abundant food access in pre-puberty predicted higher all-cause mortality in his grandsons, but not granddaughters, driven largely by cancer deaths. It did not reproduce the original findings for diabetes or cardiovascular mortality (Vågerö et al., Nature Communications, 2018).
The Father's Sperm. Fathers contribute more than DNA. In mice fed a high-fat diet, specific small RNAs in sperm changed. When researchers injected those sperm RNA fragments into normal embryos, the offspring developed metabolic disorders (Chen et al., Science, 2016). In humans, paternal overweight at conception has been associated with a doubled risk of offspring obesity (Tomar et al., Nature, 2024). Encouragingly, in one mouse study the effects of a short paternal high-fat diet were transient and disappeared after four weeks on a normal diet (Alipoor et al., Journal of Internal Medicine, 2025).
Multigenerational BPA Effects. Beyond the six-generation study, maternal BPA exposure at levels representative of human exposure produced higher body fat and impaired glucose control in both children and grandchildren of exposed mice, linked to inherited methylation changes at the Igf2 gene (Susiarjo et al., Endocrinology, 2015).
Your Fat Cells Have a Memory
If you have ever lost weight and watched it return, this finding matters.
Researchers analyzed fat tissue from people before and two years after bariatric surgery. Even after substantial weight loss, fat cells, blood vessel cells, and precursor cells retained transcriptional changes associated with obesity. In mice, obesity left persistent epigenetic changes in fat cells that impaired their function, and mice carrying this "obesogenic memory" regained weight faster when re-exposed to a high-fat diet (Hinte et al., Nature, 2024). In those mice, promoters that should have been active remained silenced and the enhancer landscape stayed remodeled (Greenhill, Nature Reviews Endocrinology, 2025).
This may help explain the frustrating "yo-yo" effect. Weight regain is not simply a failure of willpower. Your body may be responding to lasting biological changes from its earlier state.
The Honest Part: Cause or Footprint?
Here is what many discussions of epigenetics leave out. A mark that appears alongside obesity is not necessarily a mark that causes it.
In a large epigenome-wide study, researchers identified 187 methylation sites associated with BMI. When they tested the direction of influence, methylation at most sites was a consequence of BMI. Only one site, near the NFATC2IP gene, showed evidence of possibly driving BMI (Wahl et al., Nature, 2017). A longitudinal study following people over about six years found that baseline BMI predicted later methylation changes, while no methylation site predicted later BMI (Sun et al., Circulation Research, 2019). Similar results have been reported in multi-ethnic Asian populations (Chen et al., Clinical Epigenetics, 2021).
The exceptions matter. Methylation near NFATC2IP predicts future weight gain, and the SREBF1 site appears to have a causal effect on BMI (Ling et al., Nature Reviews Endocrinology, 2022).
There is another limitation. Most studies measure methylation in blood, which is easy to collect. But fat tissue is where much of the biology happens, and results from bulk blood or tissue samples can be confounded by differences in cell composition (Hinte et al., Nature, 2024).
The most accurate conclusion: obesity changes epigenetic marks far more often than those marks cause obesity. Most marks are downstream footprints. A small number may be genuine triggers.
Can Epigenetic Marks Be Rewritten?
Some can, at least in part.
- Diet. In women with severe obesity, a short-term reduced-calorie diet changed methylation at 16,064 sites across the genome. However, 878 sites remained different from normal-weight women after the diet (Nicoletti et al., European Journal of Clinical Nutrition, 2020). A very low calorie ketogenic diet shifted the methylome toward a pattern resembling normal-weight volunteers (Crujeiras et al., Clinical Nutrition, 2021).
- Bariatric surgery. After surgical weight loss, methylation of LINE1, POMC (an appetite-regulating gene), and LEP (the leptin gene) in the rectal lining returned to levels similar to those of people without obesity (ElGendy et al., International Journal of Obesity, 2023). In skeletal muscle, gastric bypass restored normal promoter methylation in genes involved in metabolism and mitochondrial function (Lagarde et al., Oncogene, 2024).
- Exercise. Regular physical activity has been linked to beneficial methylation changes at metabolic genes such as PPARGC1A and GLUT4 (Fazeli and Soleimani Samarkhazan, Obesity Surgery, 2025).
- Medications. Metformin, thiazolidinediones, GLP-1 receptor agonists, and SGLT2 inhibitors have all shown epigenetic remodeling effects in liver, muscle, and fat tissue (Ferder et al., Physiological Genomics, 2026). These medications are prescribed for specific clinical purposes, and their epigenetic effects are still being studied.
Not every mark returns to baseline. Researchers describe obesity-related epigenetic memory as a "graded decay" rather than absolute permanence: it fades, but it can persist after weight loss and limit full reversibility (Ferder et al., Physiological Genomics, 2026). That does not make regain inevitable. It means long-term support matters.
What This Means for You
Obesity is not simply about behavior, willpower, or laziness. Body weight is shaped by genes, environment, development, and the chemical marks that connect them. Some of those marks reflect conditions you never chose. Some simply record what your body has already experienced. And some can change with diet, activity, weight loss, and medical treatment.
That is a more complicated story than "just eat less." It is also a more honest one, and it is why effective obesity treatment, including understanding why bariatric surgery works, goes well beyond willpower.
Further Information
For additional context and a deeper professional perspective, you can read the companion LinkedIn article linked below.
Read on LinkedIn: Obesity Is Written in Your Genes, But Not How You Think
Medical Topics Covered
Dr. de la Cruz‑Muñoz specializes in advanced minimally invasive and robotic surgery, metabolic disease management, and comprehensive long‑term patient care. His work integrates clinical expertise, academic leadership, and a commitment to evidence‑based medicine. He has trained surgeons across the country, contributed to national guidelines, and is widely regarded for his outcomes, innovation, and patient‑centered approach.
Patients choose Dr. de la Cruz‑Muñoz for his combination of surgical excellence, metabolic expertise, and lifelong dedication to improving patient health through compassionate, high‑quality care.
Procedures We Offer
Conditions

FAQ
Bariatric surgery does not change your DNA sequence, but it can change the epigenetic marks that control how some of your genes behave. After surgical weight loss, methylation at genes tied to appetite and fat signaling (POMC and LEP) and at LINE1 returned to levels similar to people without obesity. The effects may even reach the next generation. Children born after their mother's bariatric surgery showed methylation differences in thousands of genes involved in blood sugar control, inflammation, and leptin signaling compared with siblings born before it. Not every mark resets, and researchers are still working out which changes matter most for long-term health.
Weight regain is not simply a failure of willpower: fat tissue appears to hold on to a molecular "memory" of obesity even after major weight loss. In human fat tissue studied two years after bariatric surgery, some gene activity patterns linked to obesity were still present. In mice, these lasting changes were traced to stable epigenetic marks in fat cells, and the mice carrying them regained weight faster when they were fed a high-fat diet again. This may help explain why keeping weight off is harder than losing it. It does not make regain inevitable, and it supports long-term follow-up, nutrition support, and medical therapy when needed after surgery.
People Also Ask
Obesity is recognized as a chronic disease by major medical organizations, including the American Medical Association. It is influenced by genetics, hormones, brain signaling, environment, sleep, stress and medications — not simply willpower. Like other chronic diseases, it often requires long-term treatment, which may include nutrition, activity, behavioral support, medications and surgery. Lifestyle matters, but biology largely determines how the body responds.
The body has powerful systems that defend its weight. When you lose weight, hunger hormones rise, fullness hormones drop, the brain becomes more focused on food and metabolism slows — a process called metabolic adaptation. Genetics, sleep, stress, medications and the modern food environment add to the challenge. These are biological forces, not character flaws, which is why medical treatment is often needed.
Genetics play a major role. Studies of twins and adopted children suggest that roughly 40–70% of the variation in body weight is inherited, influencing appetite, fullness, metabolism and where fat is stored. Many genes each contribute a small effect, and a few rare single-gene forms cause severe early obesity. Genes interact with environment, sleep, stress and medications, so a family history raises risk but does not determine destiny.
GLP-1 medications mimic a natural gut hormone released after eating. They act on the brain to reduce hunger and increase fullness, slow stomach emptying so meals last longer, quiet "food noise," and improve insulin release and blood sugar. Tirzepatide also acts on a second hormone, GIP. In clinical trials, these medications produced average weight loss of about 15–20% (Wilding et al., New England Journal of Medicine, 2021; Jastreboff et al., New England Journal of Medicine, 2022).
Yes. Obesity is associated with more than 200 health conditions, including type 2 diabetes, high blood pressure, high cholesterol, heart disease, stroke, sleep apnea, fatty liver disease, reflux, osteoarthritis, infertility and PCOS, depression and at least 13 types of cancer. Losing weight — through medical treatment or bariatric surgery — improves or resolves many of these conditions.
Dieting alone usually cannot overcome the body's biological defenses against weight loss. As weight falls, hunger hormones rise, fullness hormones fall and metabolism slows — changes that can last for years. Most people regain the majority of diet-related weight loss within several years. That is not a failure of willpower, and it is why treatments that address appetite biology, such as GLP-1 medications and bariatric surgery, are more effective for many people with obesity.
Consider medical evaluation when excess weight affects your health, mobility or quality of life, or when you have related conditions such as prediabetes, diabetes, high blood pressure, sleep apnea, fatty liver or joint pain. Guidelines support medications for adults with a BMI of 30 or higher, or 27 or higher with a weight-related condition, and surgery for a BMI of 35 or higher, or 30 or higher with metabolic disease. Early treatment can help prevent complications.
Yes. Bariatric surgery treats the biological drivers of obesity, changing hunger and fullness hormones, insulin sensitivity and the weight the body defends. It is the most effective and durable treatment for severe obesity, producing large, sustained weight loss and improving or resolving diabetes, high blood pressure, sleep apnea and fatty liver. Long-term studies show it also reduces the risk of heart disease, cancer and early death.